Monday, May 22, 2024
FDA Updates AI Medical Device Guidance: Source Code Traceability Now Required
by
Published
Views:
Federal regulators in the United States have introduced a significant procedural shift for artificial intelligence–based medical devices, with implications spanning global supply chains—particularly for manufacturers in China specializing in edge-computing hardware for clinical decision support. Though the exact effective date remains unspecified, the update was formally issued on May 11, 2024, and signals a tightening of premarket transparency expectations for AI/ML Software as a Medical Device (SaMD). The change directly affects export readiness, certification timelines, and internal development governance for firms targeting U.S. market access.
Event Overview
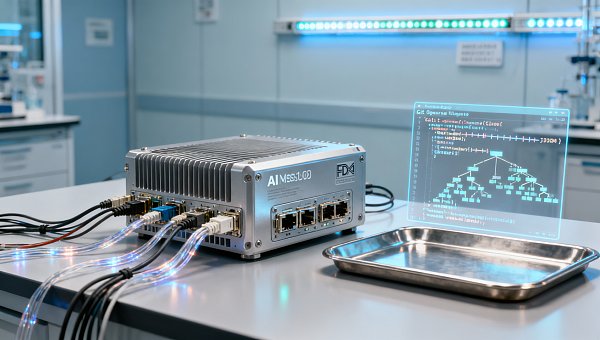
The U.S. Food and Drug Administration (FDA) updated its Artificial Intelligence/Machine Learning (AI/ML)-Based Software as a Medical Device (SaMD) Software Lifecycle Management Guideline on May 11, 2024. The revision mandates that AI-enabled diagnostic modules submitted for FDA review must include comprehensive source code version control records and a demonstrably verifiable traceability path—from initial commit through deployment and post-market updates. This requirement applies to both cloud-hosted and embedded AI functions, including those deployed on edge computing platforms used in point-of-care or intraoperative settings.
Industries Affected
Direct Exporters (e.g., SMT Precision Metrics, Smart Power Grids): These companies develop and market AI-integrated hardware modules—often deployed in radiology workstations, pathology scanners, or portable ultrasound systems. Under the new guidance, their FDA 510(k) or De Novo submissions must now include auditable source code lineage documentation, not merely model weights or inference logs. This extends submission preparation time and increases reliance on formal DevSecOps toolchains.
Raw Materials & Component Suppliers: Firms supplying FPGA SoCs, low-power AI accelerators (e.g., NPU chips), or certified memory modules face indirect but growing pressure. While not directly regulated by FDA, their component-level documentation—especially regarding firmware update mechanisms and boot-time integrity verification—must now align with downstream traceability requirements. Buyers are increasingly requesting IEC 62304-compliant supplier declarations.
Contract Manufacturing & Hardware Integration Firms (e.g., Shenzhen- and Suzhou-based EMS providers): These entities often handle final assembly, firmware flashing, and calibration of AI-enabled medical hardware. The updated guidance raises expectations for their quality management systems: firmware signing keys, build environment provenance, and immutable audit logs for every production batch are now part of FDA’s evidentiary threshold—not just for the OEM, but for contract partners involved in software-integrated manufacturing steps.
Regulatory & Certification Service Providers: Third-party notified bodies and FDA regulatory consultants must adapt assessment protocols. ISO/IEC 23053:2025 (a newly published standard for AI system documentation in health) is now routinely paired with IEC 62304 during gap analyses. Demand has surged for auditors trained in Git-based code review workflows and SBOM (Software Bill of Materials) validation—skills previously uncommon in traditional medical device QA teams.
Key Considerations and Recommended Actions
Adopt dual-standard development governance
Companies should align internal SDLC practices with both IEC 62304 (for safety-critical software lifecycle) and ISO/IEC 23053:2025 (for AI-specific documentation rigor). This includes maintaining immutable commit histories, signed build artifacts, and traceable mappings between training data versions, model checkpoints, and deployed binaries.
Reassess supplier qualification criteria
OEMs must now require documented evidence from component vendors—including secure boot attestation logs, firmware update rollback prevention mechanisms, and version-controlled bootloader configurations—to satisfy FDA’s end-to-end traceability expectation.
Integrate traceability into CI/CD pipelines—not just documentation
Traceability is no longer a static appendix. It must be machine-verifiable: automated generation of SBOMs, cryptographic linking of code commits to FDA submission packages, and real-time dashboards showing version alignment across development, test, and production environments are becoming baseline expectations.
Editorial Perspective / Industry Observation
Observably, this update marks a structural pivot—not merely a procedural tweak. The FDA is shifting from evaluating AI outputs (e.g., sensitivity/specificity metrics) toward validating the reproducibility and controllability of the entire development process. Analysis shows this reflects broader international convergence: the EU’s MDR Annex XVI and Canada’s Health Canada Guidance on AI SaMD similarly emphasize lifecycle transparency, though the FDA’s explicit linkage to source code versioning is currently the most granular. From an industry perspective, this is better understood as a signal that regulatory acceptance is now contingent on engineering discipline—not just clinical validation.
Conclusion
This policy update does not raise the bar for clinical performance—but it fundamentally redefines what constitutes acceptable evidence of reliability. For global AI medical hardware developers, the implication is clear: traceability is no longer optional infrastructure; it is a core product requirement. The long-term impact may accelerate consolidation among firms with mature DevOps maturity—and widen the compliance gap for those still relying on manual release tracking or fragmented toolchains.
Source Attribution
U.S. FDA, Artificial Intelligence/Machine Learning (AI/ML)-Based Software as a Medical Device (SaMD) Software Lifecycle Management Guideline, Revision dated May 11, 2024. Available at: fda.gov/samd-ai-ml-guidance.
ISO/IEC 23053:2025, Information technology — Artificial intelligence — Framework for AI system documentation, published March 2025.
IEC 62304:2006+A1:2015, Medical device software — Software life cycle processes.
Note: Implementation timelines for certain provisions (e.g., retrospective traceability for legacy models) remain under public consultation; stakeholders should monitor FDA’s docket FDA-2023-N-1829 for updates.
The Archive Newsletter
Critical industrial intelligence delivered every Tuesday. Peer-reviewed summaries of the week's most impactful logistics and market shifts.